Thermodynamic Potentials and Maxwell Relations
From the archive (2024). Corrected in October 2026; see the note at the end.
Below we derive the four thermodynamic potentials and give a short derivation of a Maxwell relation.
Throughout, the system is closed: no matter is exchanged with the surroundings. For an open system we would also need the chemical potential.
The fundamental relation
The first law of thermodynamics reads \(dU = \delta Q - \delta W\). For quasi-static work, \(\delta W = P\,dV\), and for a reversible process \(dS = \delta Q / T\), so \(\delta Q = T\,dS\). Combining these gives the fundamental thermodynamic relation
\[dU=T\,dS-P\,dV .\]Although we used a reversible process to derive it, it relates state functions only, so it holds for any change between equilibrium states.
Enthalpy
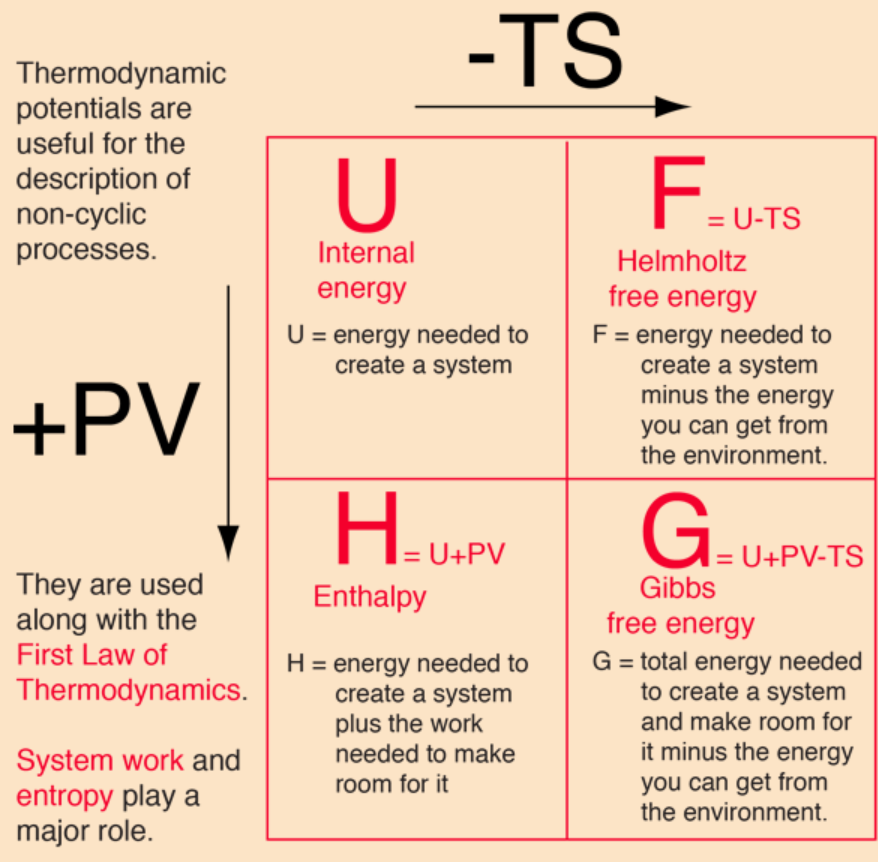
The enthalpy is the natural potential when the entropy and pressure are held fixed (\(dS=0\), \(dP=0\)): it is minimised at equilibrium under those conditions. To trade the \(dV\) in \(dU\) for a \(dP\), define
\[H=U+PV .\]Then
\[dH=dU+d(PV) =T\,dS-P\,dV+P\,dV+V\,dP =T\,dS+V\,dP .\]Helmholtz free energy
The Helmholtz free energy is minimised in processes at constant temperature and volume (\(dT=0\), \(dV=0\)). It is defined by
\[F=U-TS,\]and you can check that \(dF = -S\,dT - P\,dV\).
Gibbs free energy
The Gibbs free energy is minimised in processes at constant temperature and pressure (\(dT=0\), \(dP=0\)), the conditions of most laboratory chemistry. It is defined by
\[G=U+PV-TS=H-TS,\]and you can check that \(dG=V\,dP-S\,dT\).
A way to remember them
I struggled to memorise these relationships until I came across this diagram:
An example of a Maxwell relation
For a function \(f(x^1, \ldots, x^n)\),
\[df = \sum_{i} \frac{\partial f}{\partial x^i}\, dx^i .\]Comparing with \(dG = -S\,dT + V\,dP\), we read off
\[\left( \frac{\partial G}{\partial T} \right)_P = -S, \qquad \left( \frac{\partial G}{\partial P} \right)_T = V .\]Now take mixed second derivatives:
\[\left(\frac{\partial}{\partial P}\left(\frac{\partial G}{\partial T}\right)_P\right)_T = -\left(\frac{\partial S}{\partial P}\right)_T, \qquad \left(\frac{\partial}{\partial T}\left(\frac{\partial G}{\partial P}\right)_T\right)_P = \left(\frac{\partial V}{\partial T}\right)_P .\]By the symmetry of second derivatives, the two are equal, giving the Maxwell relation
\[\left(\frac{\partial S}{\partial P}\right)_T = -\left(\frac{\partial V}{\partial T}\right)_P .\]This is useful because changes in entropy are hard to measure directly, whereas thermal expansion is easy to measure. Each of the other three potentials gives a Maxwell relation in the same way; deriving them is a good exercise.
Changes on 25 October 2026
What was wrong. Heat and work were written as exact differentials, \(dQ\) and \(dW\); they are inexact, \(\delta Q\) and \(\delta W\). The Maxwell relation also did not say which variable is held fixed in the outer derivative.
What changed. Both are fixed, and the Helmholtz free energy, previously left as an exercise, is derived.